
A collaborative team of researchers from the California Institute of Technology (Caltech) and Yale University has achieved a major milestone in theoretical physics by developing a computational method capable of precisely calculating the Kondo effect in real, physical materials. For decades, the scientific community has been forced to rely on simplified theoretical approximations that could only capture the complex quantum phenomenon roughly. By bypassing these traditional approximations and working directly from a material’s authentic atomic and electronic architecture, this newly published research opens unprecedented pathways for the computer simulation of advanced quantum systems, including high-temperature superconductors.
The findings, detailing this quantum chemistry breakthrough, were formally published in the prestigious journal Science. The research effort was spearheaded by lead authors Linqing Peng, a 2025 PhD graduate, and Tianyu Zhu of Yale University. Both scholars initiated and developed the foundational aspects of the project while working within the laboratory of Garnet Chan, the Bren Professor of Chemistry and director of the Rudolph A. Marcus Center for Theoretical Chemistry at Caltech.
The Historical Context of the Kondo Effect
To understand the magnitude of this breakthrough, one must examine the historical landscape of condensed matter physics. In standard semiconductor materials like silicon, which form the bedrock of modern microelectronics and consumer technology, interactions among electrons are relatively weak. Because these interactions are minimal, physicists can frequently ignore them when mapping out the overall electrical and thermal behavior of the bulk material.
However, this traditional framework fails entirely when applied to strongly correlated materials—a class of substances vital for next-generation quantum technologies. In strongly correlated systems, the behavior of a single electron is inextricably tied to the dynamic states of all nearby electrons. Calculating these interconnected trajectories creates what physicists call a many-body problem, a mathematical nightmare where traditional analytical tools quickly break down.
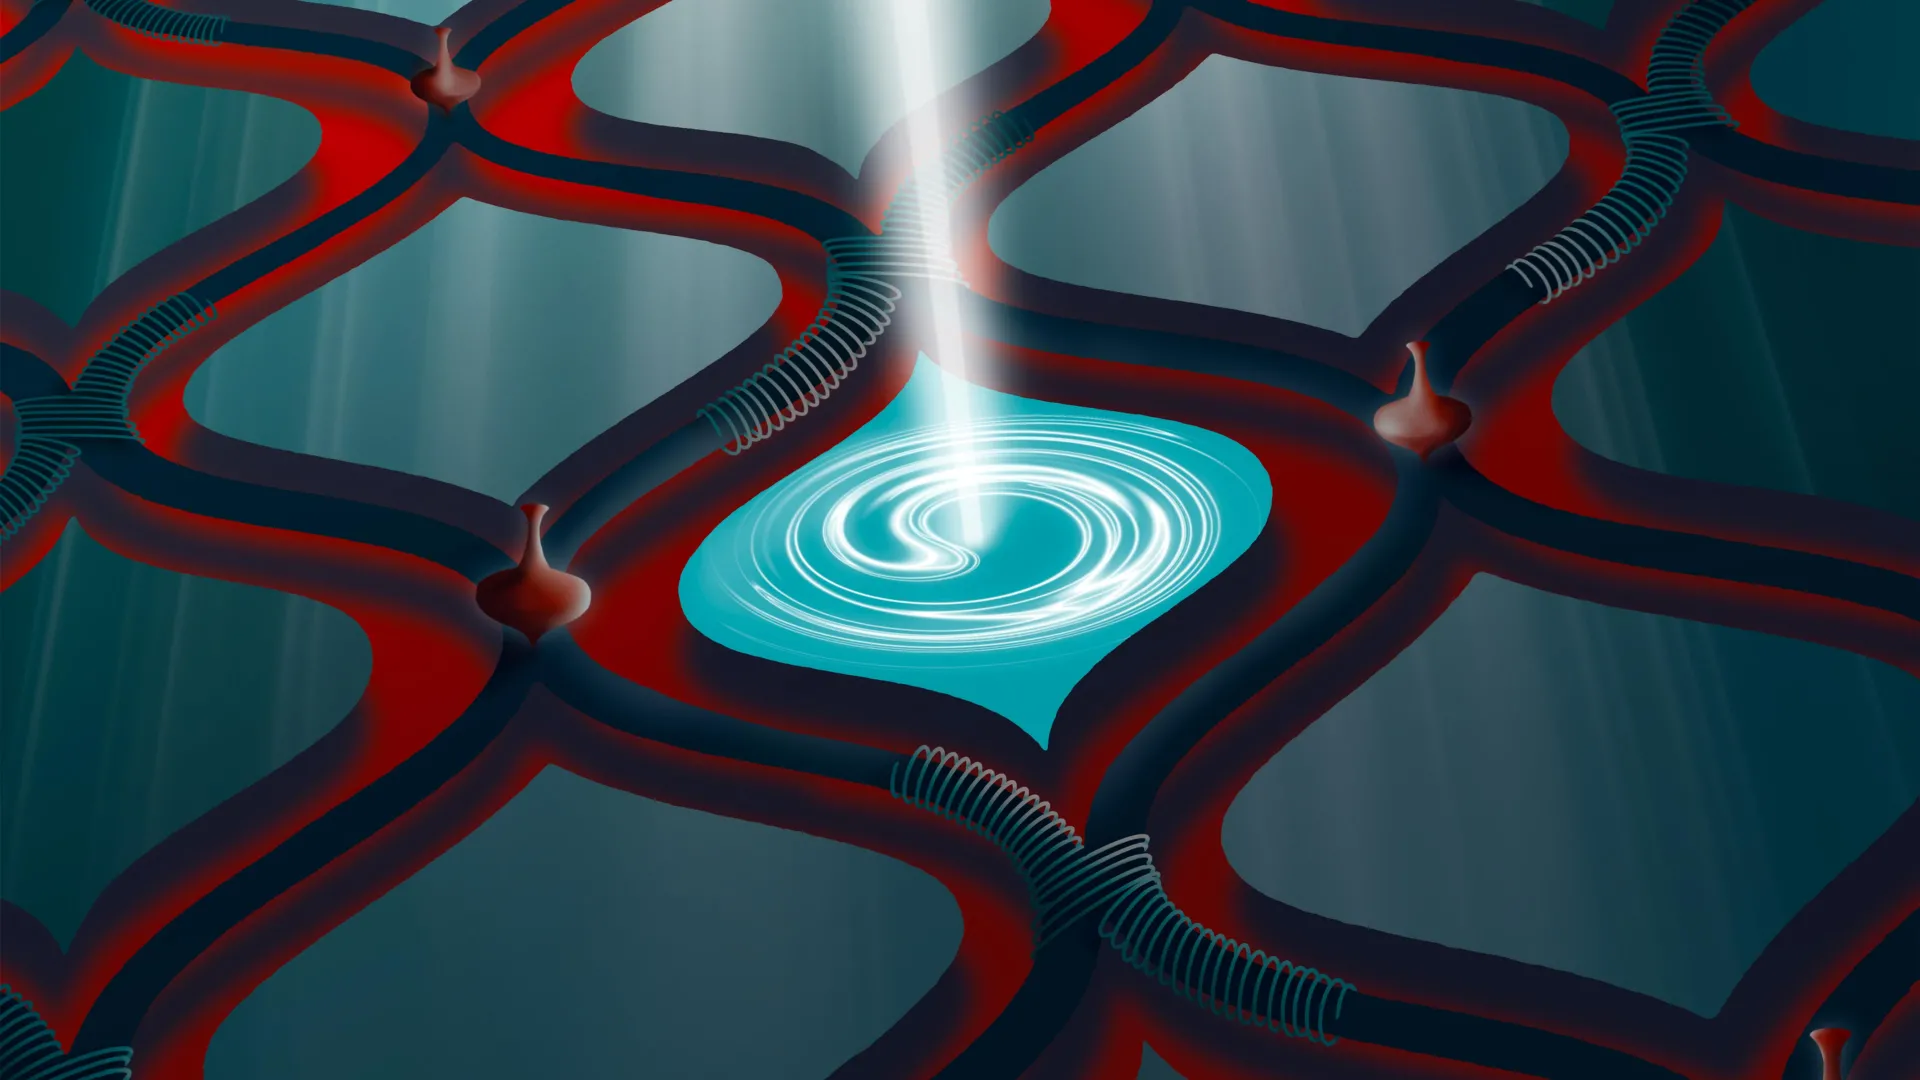
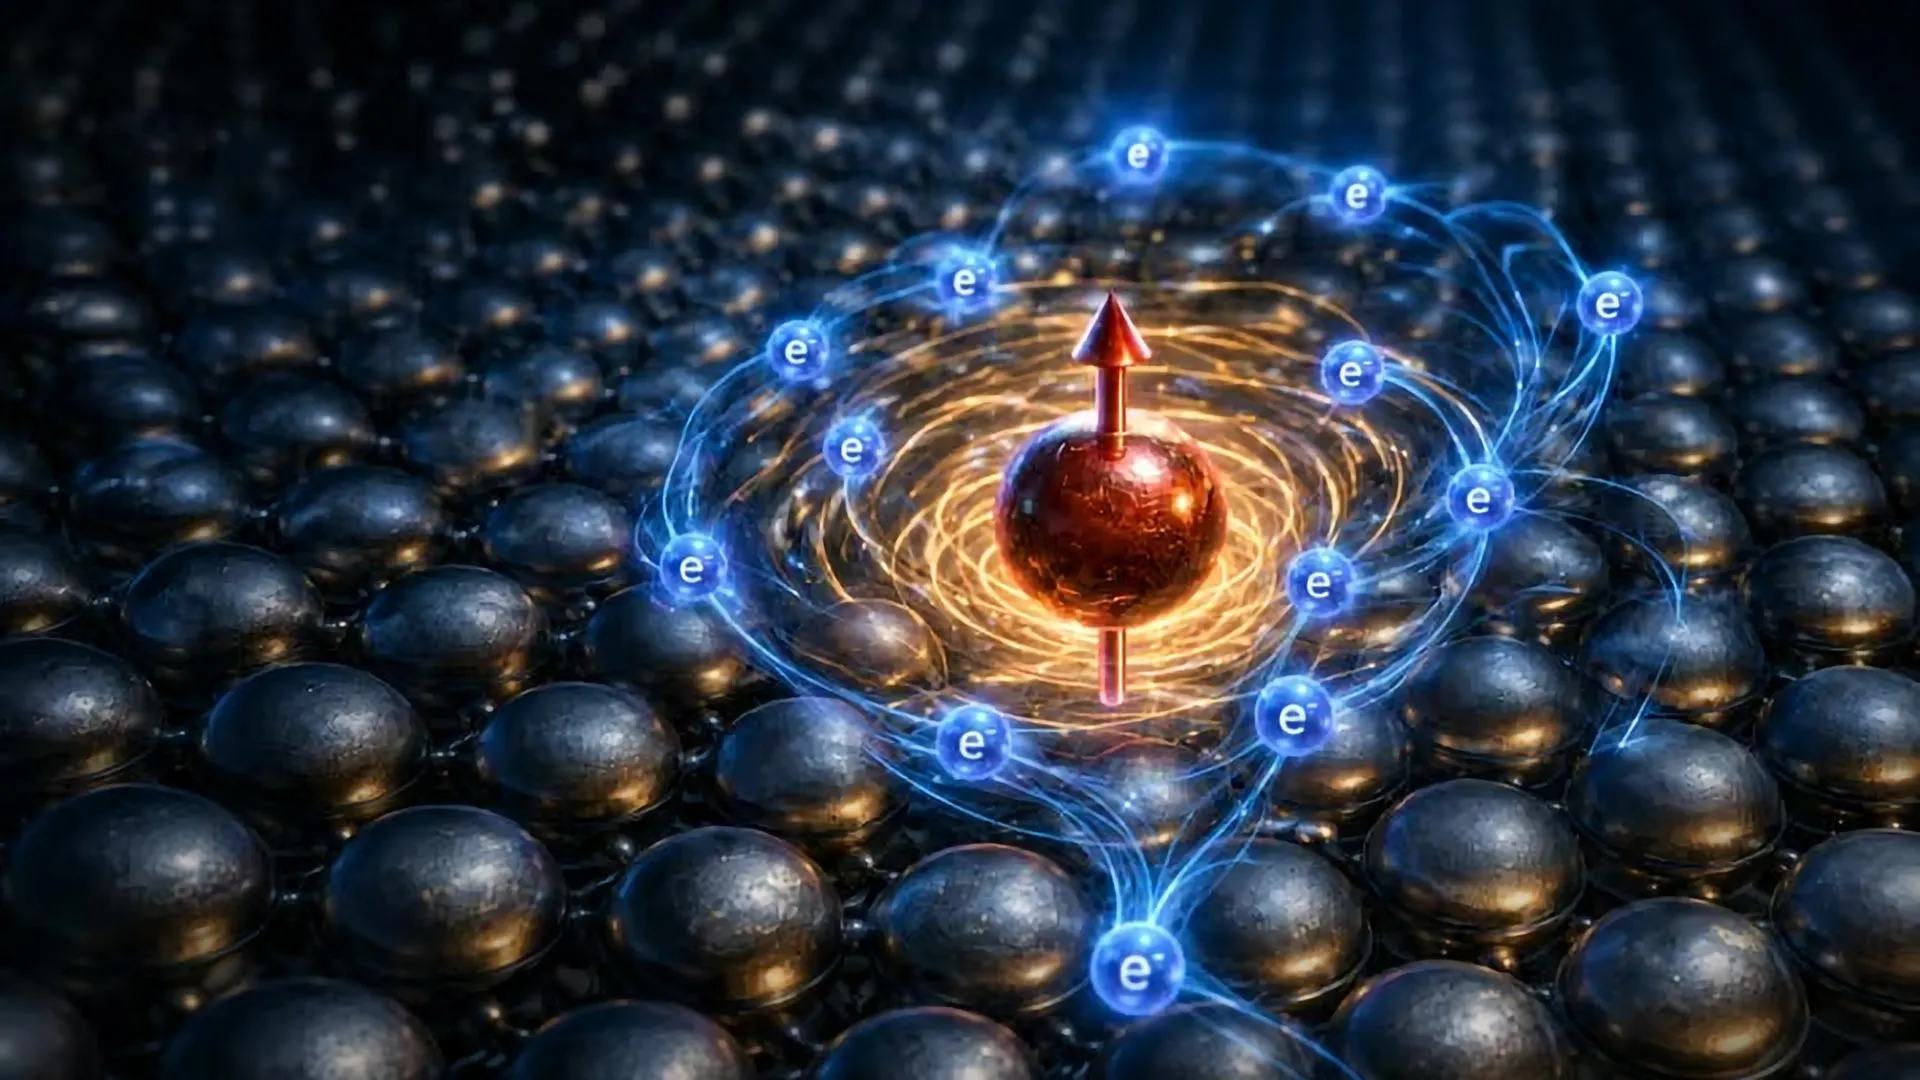
The Kondo effect stands as a textbook example of a strongly correlated many-body challenge. Discovered experimentally decades ago, the phenomenon occurs when a minute quantity of a magnetic impurity, such as an iron or manganese atom, is embedded within a non-magnetic metal lattice like copper. Under normal conditions, lowering the temperature of a pure metal causes its electrical resistance to drop steadily, facilitating smoother current flow as thermal scattering diminishes.
Yet, when a magnetic impurity is introduced, the electrical resistance defies conventional expectation. As the material cools, the resistance drops to a specific minimum before reversing trajectory and climbing steadily upward as temperatures approach absolute zero. This anomalous resistance minimum is the definitive signature of the Kondo effect, driven by interactions between the localized magnetic moment of the impurity and the collective pool of conduction electrons flowing past it.
A Chronology of Theoretical Progress
The general theoretical framework explaining the Kondo effect was largely mapped out during the 1970s. A pivotal contribution came from Caltech alumnus Kenneth Wilson (PhD ’61), whose pioneering work on the renormalization group earned him the Nobel Prize in Physics in 1982. Wilson’s work provided a mathematical language to understand how electrons at vastly different energy scales interact near the impurity.
Despite this theoretical foundation, calculating the precise resistance curves and exact Kondo temperatures for specific, real-world elemental combinations remained out of reach. Traditional physics approaches tackled the problem by stripping away real-world chemical complexity. Researchers would reduce a material’s electronic structure to a handful of idealized orbitals—mathematical zones around an atomic nucleus where electrons reside—and feed those simplified proxies into generalized model Hamiltonians, such as the Anderson impurity model or the Kondo model.
While these models provided profound qualitative insights, they routinely fell short when asked to make quantitative predictions for actual, physical materials containing specific transition-metal atoms. The gap between abstract mathematical models and messy, real-world chemistry persisted as a major bottleneck in materials science for over forty years.
A Quantum Chemistry Paradigm Shift
To bridge this decades-old chasm, the Caltech-Yale team abandoned conventional model-based reductions and adopted a radically different strategy. They took highly accurate computational algorithms and algorithmic toolkits originally engineered in the field of quantum chemistry to study complex molecular structures, and repurposed them to simulate solid-state quantum materials.
By treating the magnetic impurity and its surrounding metallic environment through the lens of advanced quantum chemistry, the researchers were able to simulate the system with near-exact fidelity. This methodology allowed them to retain the complete chemical complexity of the atomic interactions without resorting to the drastic simplifications that plagued earlier generations of researchers.
The research team rigorously tested their novel computational framework on a series of experimental benchmarks: seven distinct transition-metal atoms embedded directly into a copper host lattice. When compared against established model-based predictions, the new computational approach yielded results that were up to two orders of magnitude more accurate, marking a quantitative leap forward in predictive solid-state physics.
Official Responses and Academic Perspectives
Reflecting on the implications of the work, senior author Garnet Chan emphasized the foundational nature of the achievement.
"It is now possible to predict the properties of some complicated materials purely through computation without referring to experiment," said Chan, who also serves as a Simons Investigator in Physics. "These first materials that we have studied are like a baby step, or a prototype problem, along the way to more complex phenomena such as high-temperature superconductors and quantum magnets."
Lead author Linqing Peng echoed these sentiments, highlighting the broader movement within computational physics toward first-principles material design.
"We are in an exciting era in which faithful predictive quantum descriptions of the full chemical complexity of real materials are coming within reach," Peng noted. "It is becoming realistic to predict material-specific behavior of correlated electrons from first principles, even in some of the most challenging classes of quantum materials."
Peng further emphasized how this methodology could transform experimental physics. "This is an important step toward computationally designing materials whose functions emerge from intricate correlated physics, such as high-temperature superconductivity, where the large chemical space and competition among many phases call for predictive theory to help focus the experimental search for new materials. I am excited to see what new materials breakthroughs this theory will enable in the future."
In addition to Peng and Zhu, the published study, titled Toward an exact quantum many-body treatment of Kondo correlation in magnetic impurities, lists several key contributors from the Caltech ecosystem. These include former postdoctoral scholar Huanchen Zhai, current postdoctoral scholar Runze Chi, and Zhi-Hao Cui (PhD ’23), who completed a significant portion of the foundational calculations as a graduate student.
Broader Impact and Future Implications
The publication of this methodology arrives at a critical juncture for both materials science and the technology sector. As the global scientific community races to develop stable quantum computers, ultra-efficient power grids, and zero-loss energy transmission systems, understanding strongly correlated quantum materials has transformed from an academic pursuit into an urgent technological necessity.
High-temperature superconductors—materials capable of conducting electricity with zero electrical resistance at temperatures vastly higher than conventional superconductors—remain one of the holy grails of modern physics. However, their internal mechanics are governed by intense electron correlations that defy traditional computational methods. By proving that complex many-body phenomena like the Kondo effect can be calculated directly from a material’s true atomic structure, the Caltech and Yale team has provided researchers with a powerful new computational compass.
Furthermore, this development signals a broader philosophical shift in how scientific discoveries may be accelerated in the coming decades. Rather than relying solely on the slow, expensive process of trial-and-error laboratory synthesis, materials scientists may increasingly rely on high-fidelity quantum simulations to screen thousands of candidate materials digitally before synthesizing them in a lab.
Financial and institutional backing for the research was provided by several key federal agencies, reflecting the strategic importance of the work to national scientific infrastructure. Support came from the Air Force Office of Scientific Research via its Multidisciplinary University Research Initiative (MURI) program, the United States Department of Energy through its Center for Molecular Magnetic Quantum Materials, and the National Science Foundation. As these computational tools continue to mature, the boundary between theoretical quantum chemistry and practical solid-state engineering will continue to dissolve, heralding a new era of predictive materials design.