
Scientists at the California Institute of Technology (Caltech) and Yale University have successfully developed a groundbreaking computational methodology to precisely calculate the Kondo effect in specific, real-world materials. This achievement eliminates a decades-long reliance on simplified, approximate theoretical models. Published in the prestigious journal Science, the research represents a major milestone in theoretical physics and quantum chemistry, bridging the gap between abstract quantum mechanics and the predictive modeling of complex chemical systems.
For generations, researchers studying strongly correlated electron systems have been forced to use generalized models that capture physical phenomena only approximately. By working directly from a material’s true atomic and electronic structure, the new approach allows scientists to calculate material-specific properties purely through computation, bypassing the need for initial experimental data fitting. This breakthrough not only solves a classic many-body physics problem but also lays a critical foundation for the future computer simulation of high-temperature superconductors and quantum magnets.
The research was spearheaded by lead authors Linqing Peng and Tianyu Zhu of Yale University. Both investigators initiated the project within the laboratory of Garnet Chan, Bren Professor of Chemistry and director of the Rudolph A. Marcus Center for Theoretical Chemistry at Caltech. Additional co-authors from Caltech include former postdoctoral scholar Huanchen Zhai, current postdoctoral scholar Runze Chi, and Zhi-Hao Cui, who completed the research as part of his doctoral studies. Financial backing for the multi-institution project was provided by the Air Force Office of Scientific Research through the Multidisciplinary University Research Initiative program, the U.S. Department of Energy via its Center for Molecular Magnetic Quantum Materials, and the U.S. National Science Foundation.
Historical Chronology and Background of the Kondo Effect
To understand the magnitude of this computational advance, one must examine the history of the Kondo effect within theoretical physics. In standard semiconductor materials like silicon, interactions among electrons are relatively weak. These interactions can frequently be ignored when describing the bulk material’s overall electrical and thermal behavior. However, in strongly correlated materials—which are vital for emerging quantum technologies—electrons constantly interact, scatter, and influence one another in complex ways.
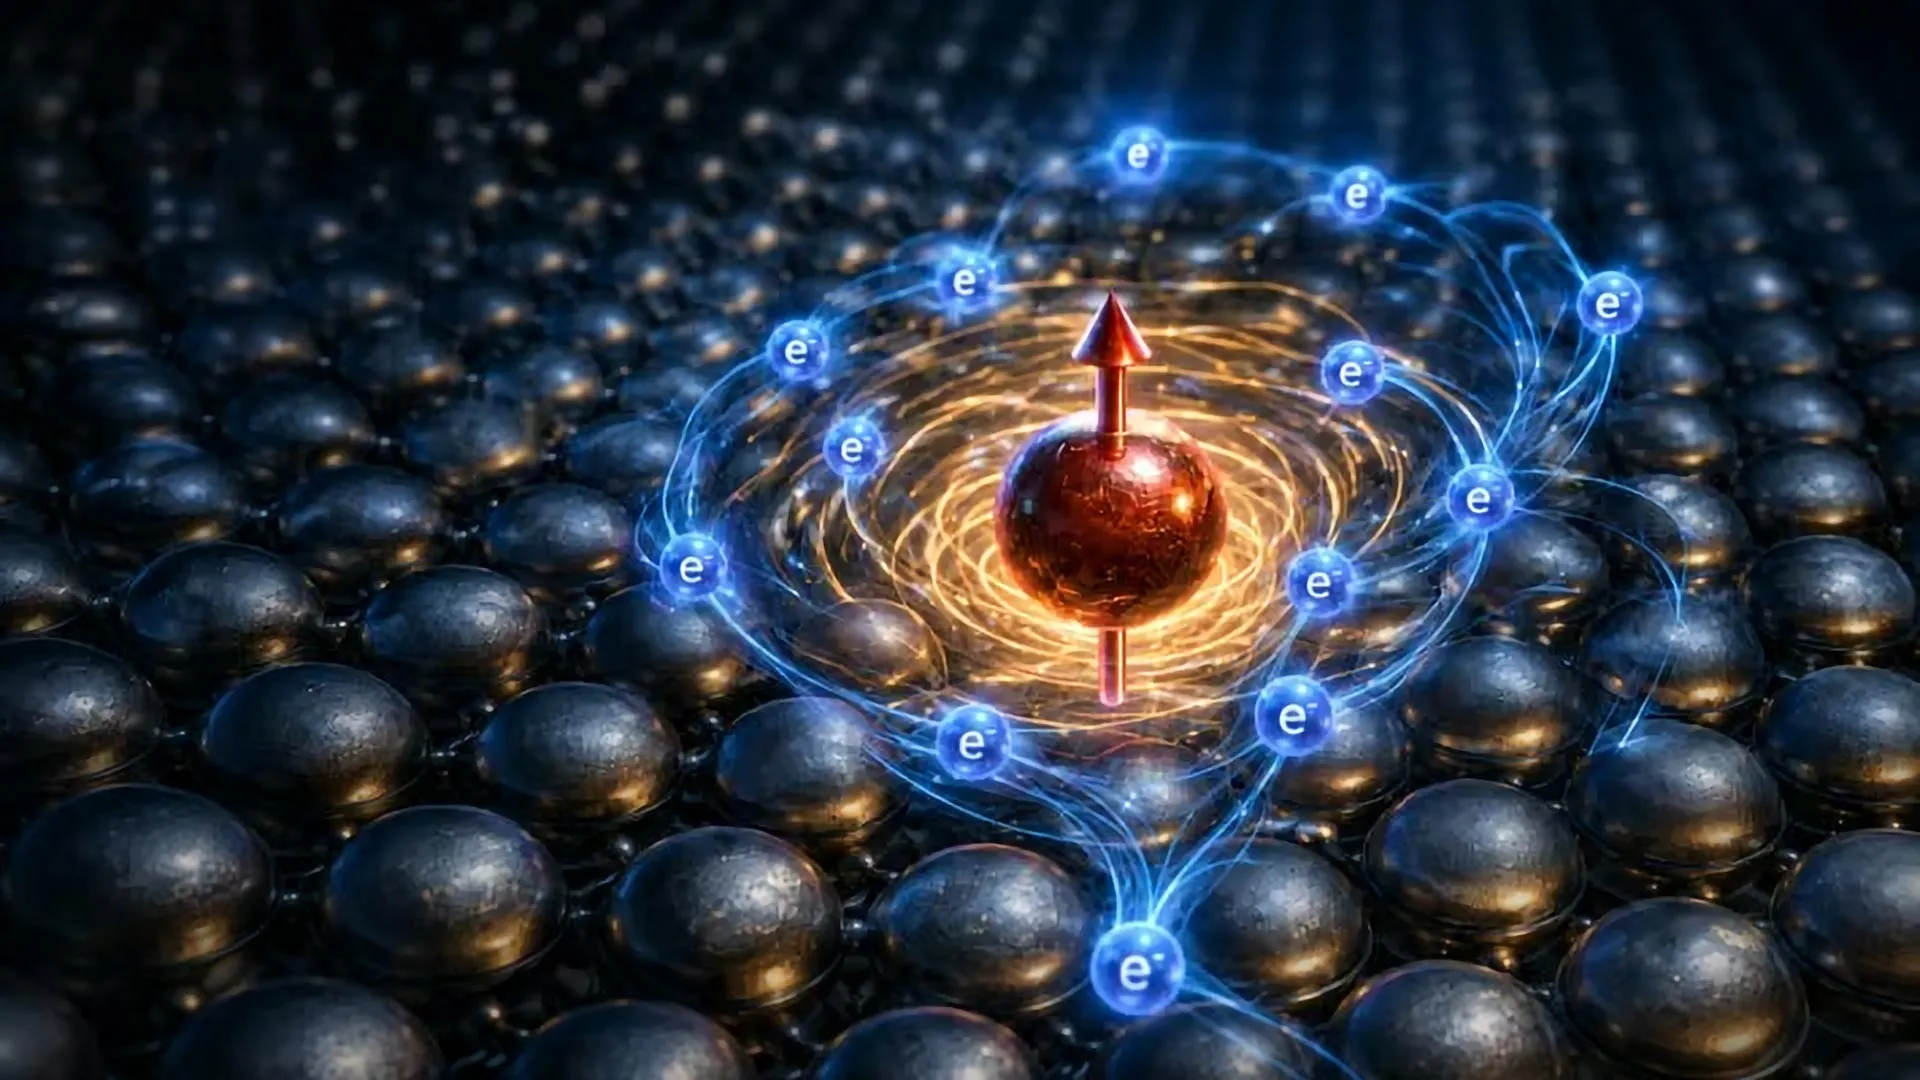
The Kondo effect serves as a classic, idealized manifestation of a strongly correlated many-body system. The phenomenon typically occurs when a solitary magnetic transition-metal atom, such as iron or manganese, is embedded as an impurity within a non-magnetic metal matrix, such as copper. Under normal thermodynamic conditions, cooling a pure metal causes its electrical resistance to decrease steadily, allowing electric current to flow with progressively less resistance.
However, when a magnetic impurity is present, this standard behavior breaks down. As the material is cooled below a specific threshold known as the Kondo temperature, the electrical resistance halts its decline, hits a local minimum, and paradoxically begins to rise again as the temperature approaches absolute zero.
This anomalous resistance minimum puzzled experimental physicists for decades until Japanese physicist Jun Kondo formulated a theoretical explanation in 1964 using perturbation theory. The phenomenon was later fully solved in the 1970s using the renormalization group technique pioneered by Caltech alumnus Kenneth Wilson, who was subsequently awarded the Nobel Prize in Physics in 1982.
The underlying physics involves the spin of unpaired electrons within the magnetic impurity. At elevated temperatures, the impurity’s magnetic moment fluctuates freely. As the system cools below the Kondo temperature, electrons traveling through the bulk metal begin to interact strongly with the impurity’s spin. These conduction electrons can flip their own spins, partially canceling the magnetic moment of the embedded atom.
The resulting scattering events create the characteristic upturn in electrical resistance. As more electrons participate, they collectively form a quantum mechanical cloud that "screens" or completely masks the magnetism of the impurity. While physicists developed a general theoretical framework for this mechanism decades ago, applying it quantitatively to real, chemically complex materials has remained an insurmountable hurdle until now.
Methodology: Adapting Quantum Chemistry Tools
Until this recent advance, researchers relied heavily on simplified quantum models. Traditional techniques typically reduce the electronic structure of a material to a small, tractable set of atomic orbitals—regions around an atomic nucleus where electrons are most likely to be located—and apply phenomenological parameters to match experimental observations. While useful, these models sacrifice chemical realism and cannot reliably predict quantitative outcomes for specific material compositions.
The research team led by Chan and Zhu adopted an entirely different strategic paradigm. They took highly accurate computational algorithms and quantum-chemical tools originally engineered for describing the intricate wavefunctions of complex molecules and successfully adapted them to solid-state quantum materials.
This cross-disciplinary adaptation allowed the researchers to treat magnetic impurities within a metal lattice with molecular-level precision, preserving the full complexity of inter-electronic interactions. Instead of reducing the system to abstract parameters, the computation retains the exact atomic coordinates, orbital symmetries, and electronic configurations of the real-world substances under investigation.
Quantitative Results and Verification Data
To validate their new methodology, the research team tested the framework on seven distinct transition-metal atoms embedded as impurities within a copper host lattice. The performance metrics of the computation exceeded prior benchmarks by a wide margin.
According to the findings, the new computational approach yielded predictions that were up to two orders of magnitude—or roughly 100 times—more accurate than those obtained from conventional model-based techniques. For the first time, researchers could accurately compute the exact temperature at which electrical resistance transitions from falling to rising, as well as the magnitude of the resistance anomaly for specific elemental impurities, without relying on adjustable parameters derived from prior experiments.
Broader Impact and Implications for Quantum Technologies
The successful calculation of the Kondo effect in real materials marks the crossing of an important technological and theoretical threshold. In many ways, the study of isolated magnetic impurities in a metallic host serves as a foundational prototype for understanding much larger classes of correlated electron systems.
Experts note that the methodology directly addresses the primary bottleneck in modern computational materials science: the inability to simulate systems where individual particle behaviors cannot be decoupled. By demonstrating that quantum chemistry tools can accurately capture solid-state phenomena from first principles, the study opens new avenues for exploring high-temperature superconductivity, quantum magnetism, and heavy-fermion systems.
High-temperature superconductors, which conduct electricity with zero resistance at temperatures significantly higher than conventional superconductors, represent one of the holy grails of modern condensed matter physics. However, their internal mechanics are notoriously difficult to model because the behavior of any single electron is inextricably linked to the collective state of its neighbors. The enormous chemical space and fierce competition among various electronic phases in these materials have historically made experimental discovery a slow, trial-and-error process.
Official Responses and Expert Commentary
Reflecting on the broader implications of the published research, senior author Garnet Chan emphasized the predictive power of the new computational framework.
"It is now possible to predict the properties of some complicated materials purely through computation without referring to experiment," Chan stated. "These first materials that we have studied are like a baby step, or a prototype problem, along the way to more complex phenomena such as high-temperature superconductors and quantum magnets."
Lead author Linqing Peng echoed these sentiments, highlighting the transition into a new era of materials science driven by high-fidelity quantum simulations.
"We are in an exciting era in which faithful predictive quantum descriptions of the full chemical complexity of real materials are coming within reach," Peng observed. "It is becoming realistic to predict material-specific behavior of correlated electrons from first principles, even in some of the most challenging classes of quantum materials."
Peng further emphasized how predictive theory will alter the landscape of experimental research. "This is an important step toward computationally designing materials whose functions emerge from intricate correlated physics, such as high-temperature superconductivity, where the large chemical space and competition among many phases call for predictive theory to help focus the experimental search for new materials. I am excited to see what new materials breakthroughs this theory will enable in the future."
Future Directions in Computational Materials Design
As academic and industrial laboratories race to develop scalable quantum computers and advanced electronic devices, the demand for precise materials simulation tools has never been higher. Traditional trial-and-error synthesis in the laboratory is both time-consuming and economically costly, often limited by the human intuition of researchers exploring vast chemical databases.
By proving that first-principles quantum chemistry can accurately predict complex electronic phenomena like the Kondo effect, the Caltech and Yale collaboration has provided the scientific community with a robust new compass. Future research efforts will likely focus on scaling these computational algorithms to handle larger lattices, higher concentrations of impurities, and multi-component systems exhibiting competing quantum phases.
Ultimately, this methodological leap moves the scientific community closer to the long-sought goal of computational materials design—a paradigm where novel quantum materials with customized electrical, magnetic, and superconducting properties can be entirely engineered on a computer screen before a single physical sample is synthesized in the laboratory.